Assigning a room grade before the process itself has been classified reverses the order a cleanroom design actually depends on. The protection objective, the sterility requirement, and the contamination route of each process step determine which controls apply — not the other way around. Getting this sequence wrong is how sterile-style controls get copied into nonsterile processes, or how a nonsterile cross-contamination risk gets left unaddressed because the room grade looked adequate on paper.
Process Risk Differences to Define Before Assigning Room Grades
| Process classification question | Finding to establish | Design decision it informs |
|---|---|---|
| Product exposure | Where the product is exposed and whether each step is open or closed | Where local protection or room-level control must cover the process |
| Sterility requirement | Which steps require sterile processing and which are nonsterile | Whether sterile contamination-control duties are applicable |
| Contamination route | The contamination route relevant to each process step | Which control boundary must address that route |
| Operator intervention | Where intervention occurs in relation to product exposure | Whether the local-protection boundary must include the intervention |
| Material flow | The material route and its transfer points | Where transfer controls and room boundaries must be placed |
A process cannot be matched to equipment or a room grade until its exposure conditions are known in enough detail to describe where contamination could actually enter the product. This starts with identifying where the product is exposed to the room environment and whether each step in the process is open or closed. An open step, where the product or a critical surface is exposed directly to room air, carries a different contamination route than a closed step where a barrier or enclosure already separates the product from the surrounding environment. Treating these as equivalent because they occur in the same room ignores the mechanism that actually determines risk.
Sterility requirement is a separate question from exposure. A process can involve open handling without being a sterile process, and a closed step can still sit within a sterile process if an upstream or downstream step requires it. Confusing these two conditions leads either to over-specifying control for a nonsterile step or under-specifying it for a sterile one. The contamination route relevant to each step — whether it is airborne, surface-transfer, personnel-associated, or material-associated — further narrows which boundary needs to address it; a route that travels through material transfer calls for a different control point than one that travels through operator movement near an open product.
Operator intervention location matters independently of exposure type. Where intervention occurs away from the point of product exposure, the local-protection boundary can be drawn tighter around the process itself. Where intervention necessarily occurs at or near the exposure point, the boundary has to extend to cover that intervention, which changes what equipment configuration is adequate. Finally, material flow and its transfer points determine where transfer controls and room boundaries need to sit geographically, not just functionally — a process with a single material entry point has a different boundary-placement problem than one with multiple transfer points feeding the same open step.
None of these five questions can be answered from the room grade alone, which is why resolving them first is what allows the room grade and equipment selection that follow to be justified rather than assumed.
Applying Sterile Contamination-Control Duties to Open and Closed Steps
Where a process step is classified as sterile, the contamination-control duty attached to it does not stop at the equipment boundary. A sterile step requires a connected strategy across facilities, equipment, procedures, monitoring, and qualification — meaning the equipment alone cannot carry the sterility assurance that the process requires. This is a structural requirement of sterile processing: the duty is distributed across the system, and no single component discharges it independently of the others.
The practical difference between open and closed sterile steps is where that distributed duty concentrates. In an open sterile step, the facility and equipment controls have to actively maintain the sterile condition against the surrounding environment for as long as the exposure lasts, which means monitoring and procedural controls carry more of the ongoing burden during that exposure window. In a closed sterile step, the barrier itself carries more of that duty, and the facility-level controls shift toward protecting the integrity of the barrier rather than the open product directly. A design that treats both step types identically — applying the same airlock, barrier, or monitoring expectation regardless of whether the step is open or closed — has not yet resolved which boundary is actually doing the protective work.
This distinction also changes what qualification needs to demonstrate. For an open sterile step, qualification has to show that the combined facility-and-equipment system maintains the sterile condition under the conditions of actual exposure. For a closed sterile step, qualification has to show that the barrier itself performs as intended and that the facility controls around it are sufficient to protect the barrier’s integrity, which is a narrower but still necessary demonstration. Where a project substitutes one qualification approach for the other without confirming which step type is present, the qualification evidence will not match what the process actually requires.
The sterile duty does not scale down proportionally to how small or brief the open exposure is; it scales according to whether the step is open or closed and what route of contamination that openness creates. A short open exposure still requires the full distributed strategy appropriate to an open sterile step, because the exposure window itself — not its duration — is what defines the control burden.
Separating Nonsterile Product Protection from Cross-Contamination Control
Nonsterile processes still carry contamination risk, but the risk is not uniform across nonsterile processes and should not be assumed to default to a lighter version of sterile control. Two distinct risk categories apply, and they are not interchangeable. The first is product protection: preventing contamination from the room or operator from degrading product quality, even where sterility is not the requirement. The second is cross-contamination control: preventing one product, material, or process from contaminating another, which is a risk between processes rather than a risk between the room and a single process.
A nonsterile process can carry significant product-protection risk while carrying little cross-contamination risk, where only one product line ever occupies the space. Conversely, a nonsterile process can carry serious cross-contamination risk while the product-protection risk for any single run is comparatively simple to manage, where multiple products or materials move through shared equipment or shared room space. Designing controls around only one of these risks because the other was assumed to be covered by the same measures leaves the uncovered risk unaddressed.
This is also where the temptation to copy sterile-process expectations becomes most consequential. A Grade A/B airlock, barrier, or monitoring expectation developed for a sterile contamination-control strategy is not automatically relevant to a nonsterile process, even one with meaningful product-protection or cross-contamination risk. If that expectation is applied, it has to be documented and justified against the nonsterile process’s own risk profile — not inherited by default because the project team is more familiar with sterile precedents. Where a nonsterile process’s actual risk is cross-contamination between materials rather than product exposure to room air, the controls that matter are the ones governing material segregation, transfer, and equipment dedication or cleaning between uses — not room-grade airflow classifications borrowed from a sterile context.
The design question for a nonsterile process is therefore not “how close to sterile does this need to be” but “which of these two risk categories is actually present, and what control addresses that specific route.”
Locating Local Protection, Transfer, Filtration, and Room-System Boundaries
| Control boundary | Project condition used to locate it | What the equipment package should show |
|---|---|---|
| Local protection | Product-exposure points and operator interventions | Where local protection begins and ends around the selected process |
| Transfer control | Material route and transfer points | Where transfer control begins and ends |
| Filtration | Filtration selected for the process | Where filtration begins and ends in relation to the equipment and room systems |
| Room system | Assigned room grade and the limits of local and equipment controls | Where room-system control begins and ends relative to the equipment package |
Once the process risk profile is established, the equipment package has to show where each control boundary physically and functionally begins and ends, because these boundaries do not default to the edges of the room or the edges of a single piece of equipment. Local protection is located by reference to the product-exposure points and operator interventions identified earlier — it has to cover the specific points where the product is open to the environment and the points where an operator intervenes near that exposure, not simply the footprint of the unit in which the process occurs.
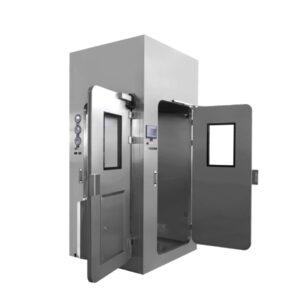
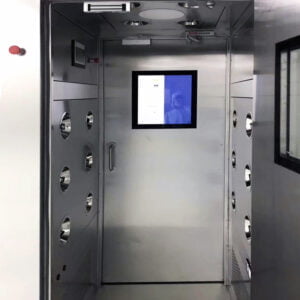
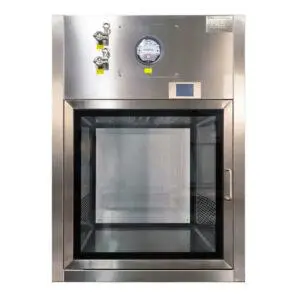
Transfer control is located differently, by reference to the material route and its transfer points rather than the product-exposure points. A transfer boundary that only addresses the primary material entry but ignores a secondary transfer point feeding the same process leaves a route open that the local-protection boundary was never designed to cover. This is where equipment families such as a pass box or VHP pass box occupy a defined role distinct from local protection: they govern the transfer boundary specifically, and their placement has to align with where the material route actually crosses between zones of differing control, not simply where room layout makes a transfer convenient.
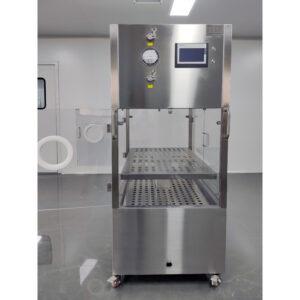
Filtration has its own boundary question, separate from both local protection and transfer: where does filtration begin and end relative to the equipment and the room system supplying air to it. A laminar airflow unit providing localized protection to a product zone performs a different filtration role than room-level filtration supplying general room air, even where both are present in the same space. Confusing the two can lead a project team to assume that room-level filtration performance substitutes for the localized protection a laminar air flow unit is intended to provide, when the two serve different exposure points entirely.
Room-system boundaries are located last, relative to the assigned room grade and the limits of whatever local and equipment controls have already been placed. The room system is responsible for the general environment outside the boundaries that local protection, transfer control, and filtration have already claimed — not for compensating where those other boundaries have been left undefined. A design review should be able to trace, for every point of product exposure and every material transfer point, which of these four boundaries is responsible, with no gap and no overlap left unexplained. The information the project team supplies about exposure points, transfer points, and the assigned room grade is what allows that boundary-by-boundary review to proceed during configuration review, rather than being reconstructed from the equipment selection after the fact.
Connecting Equipment and Facility Controls to Qualification and Monitoring
Equipment selection and facility design are not the end point of a contamination control strategy; they are inputs to a connected system that also includes procedures, monitoring, and qualification. For a sterile process, this connection is a structural requirement — the strategy has to demonstrate that facilities, equipment, procedures, monitoring, and qualification function together, not that each element independently meets its own specification. An FFU or filtration component can perform correctly in isolation and still fail to support the sterile process if the monitoring regime around it was not designed to detect the failure modes relevant to where that equipment sits in the process.
This means qualification evidence has to be read against the specific boundary the equipment occupies, established earlier. Equipment qualified only as a standalone unit does not demonstrate that the facility-equipment-procedure-monitoring combination functions as the sterile process requires; that connection has to be demonstrated at the system level, which is why qualification planning needs to reference the process’s open/closed step classification and its product-exposure points rather than treating qualification as a generic equipment checklist.
For nonsterile processes, the same connective logic applies but against a different risk basis. Monitoring for a nonsterile cross-contamination risk has to be designed around the material segregation and transfer points relevant to that risk, not around the airborne-particle monitoring points that a sterile process would specify. Where a nonsterile process’s monitoring plan is built by adapting a sterile monitoring template, the resulting plan may monitor points that are not where the nonsterile process’s actual risk route runs, while leaving the real cross-contamination or product-protection route unmonitored.
The practical implication is that monitoring and qualification planning cannot be finalized from equipment specifications alone. They depend on the same process classification — exposure points, sterility requirement, contamination route, intervention location, material flow — that determined the control boundaries in the first place. Where that classification changes during a project, for instance because a step originally assumed closed is redesigned as open, the monitoring and qualification plan built around the original classification no longer matches the process and needs to be reassessed against the new boundary.
Design Evidence That Justifies Each Sterile or Nonsterile Control
| Evidence source | Supported use in design justification | Evidence boundary |
|---|---|---|
| EU GMP Annex 1 | Connect facilities, equipment, procedures, monitoring, and qualification within a contamination control strategy for sterile medicinal products | Application to nonsterile products must be documented and justified |
| ISO 14644-4:2022 | Support cleanroom requirements through design, construction, start-up, and verification | It does not prescribe a process technology or pharmaceutical grade |
| ICH Q9(R1) | Support systematic assessment, control, communication, and review of pharmaceutical quality risks | It is a risk framework and does not set cleanroom or equipment values |
Each design decision made through this process — classifying the process, applying or withholding sterile contamination-control duties, separating product-protection from cross-contamination risk, and locating control boundaries — needs evidence to justify it, and that evidence has to match the scope of the decision it supports. EU GMP Annex 1 supports the facility-wide contamination control strategy connecting facilities, equipment, procedures, monitoring, and qualification specifically for sterile medicinal products; using it to justify a control applied to a nonsterile process requires that the application be documented and justified on its own terms, since Annex 1’s scope does not extend automatically to nonsterile products.
ISO 14644-4:2022 supports a different part of the justification: cleanroom requirements through design, construction, start-up, and verification. This standard addresses how a cleanroom is built and verified, but it does not prescribe which process technology applies or which pharmaceutical grade a given process requires — that determination has to come from the process classification work done earlier, not from the construction standard itself. A project team citing ISO 14644-4 as justification for a specific room grade has gone beyond what that standard supports; it justifies the design-construction-verification pathway, not the grade assignment.
ICH Q9(R1) supports the risk-assessment framework underlying these decisions — systematic assessment, control, communication, and review of pharmaceutical quality risks — but it does not set cleanroom values or equipment specifications itself. It is the method by which a project team can structure the classification and boundary-location decisions described above, not a source of the specific values that result from applying that method.
Matching each decision to the evidence that actually supports it means a sterile contamination-control strategy decision cites Annex 1 within its stated scope, a room design-and-verification decision cites ISO 14644-4 within its construction-and-verification scope, and a risk-assessment-process decision cites ICH Q9(R1) as the framework rather than the outcome. Where a project’s documentation cites one of these sources for a conclusion outside its stated scope — an Annex 1 citation used to justify a nonsterile control without separate documentation, for instance, or an ISO 14644-4 citation used to justify a specific grade assignment — that gap is exactly what a design review should identify before the equipment package, including any pharmaceutical modular cleanroom, pass box, or laminar air flow unit configuration proposed for the project, is treated as validated against the process risk it is meant to address.
Frequently Asked Questions
Q: Can sterile and nonsterile processes use the same cleanroom design approach?
A: Only when the controls are justified for each process. Classify product exposure, sterility requirements, contamination routes, operator interventions, and material flow before assigning room grades or deciding where local protection and room-level controls are needed.
Q: When should sterile contamination-control expectations be applied to a nonsterile area?
A: Apply a specific sterile control only when its relevance to the nonsterile product or cross-contamination risk is documented. Do not automatically carry Grade A/B airlock, barrier, or monitoring expectations into a nonsterile process without that justification.
Q: What project information should be prepared before comparing cleanroom and equipment options?
A: Prepare a process map showing open and closed steps, product-exposure points, operator interventions, contamination routes, material paths, and transfer points. This information helps define where a modular cleanroom, local airflow protection, pass-through equipment, filtration, or a room system must begin and end.
Q: How can a buyer prevent gaps between local equipment controls and the room system?
A: Require the equipment package to mark the boundaries of local protection, transfer controls, filtration, and room-system control for the selected process. Then connect those boundaries to the relevant facility provisions, procedures, monitoring, and qualification activities.
Q: What evidence is needed to justify the final control strategy?
A: Link each control to a defined process risk and use each authority only within its scope. EU GMP Annex 1 supports a sterile contamination control strategy, ISO 14644-4 supports cleanroom design through verification, and ICH Q9(R1) supports the quality-risk process; none alone supplies project-specific cleanroom grades or equipment values.